A process validation SOP defines how a site builds documented evidence that a manufacturing process can operate consistently and deliver product that meets its approved requirements. This GMP procedure covers lifecycle planning, process qualification, continued process verification, change control and revalidation. Use it as a controlled-document starting point and tailor it to the product, process, facility and applicable market.
1. Purpose and scope
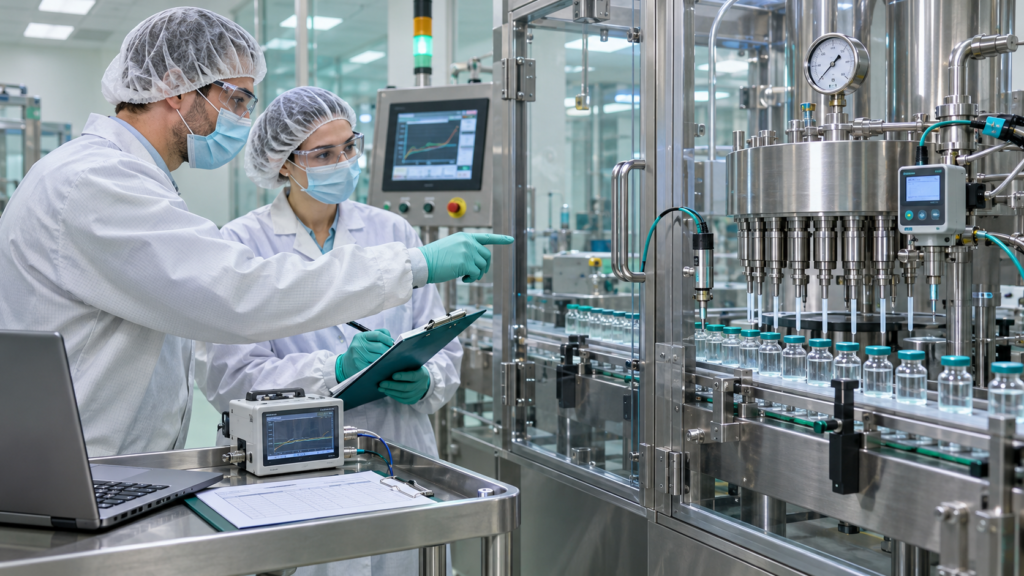
This procedure establishes responsibilities and controls for pharmaceutical process validation from process design through routine commercial manufacture. It applies to new products and processes, technology transfer, scale-up, significant process changes and ongoing monitoring. It may be used alongside equipment, facility, utility, cleaning and analytical procedure qualification; those activities require their own approved plans where applicable.
The site validation master plan or equivalent quality-system document should identify the validation strategy, governance, facilities, systems, products, responsibilities and documentation pathway. This SOP does not replace product-specific regulatory commitments, marketing-authorisation requirements or approved site procedures.
2. Regulatory and scientific principles
Apply a documented, science- and risk-based lifecycle approach. FDA guidance describes process design, process qualification and continued process verification as connected stages. EU GMP Annex 15 sets expectations for qualification and validation, while ICH quality guidelines support quality risk management and a pharmaceutical quality system. The organisation should identify which requirements apply to the product and market, then document its rationale.
Do not assume that a fixed number of batches proves validation. The protocol should justify the number, scale and design of qualification batches using process knowledge, risk, variability, complexity, prior experience and applicable regulatory commitments. Historical data can inform knowledge and risk assessment, but retrospective review alone does not replace appropriate prospective evidence for a new or changed process.
3. Responsibilities
- Quality Assurance (QA): approve the strategy, risk assessment, protocols, deviations, reports and change controls; confirm that acceptance criteria and quality-system requirements are met before approval.
- Validation or process owner: prepare the validation plan, coordinate technical inputs, assess process knowledge, define monitoring and compile the report.
- Production: execute approved instructions, record actual activities and parameters contemporaneously, and promptly report deviations or unexpected results.
- Quality Control (QC): approve sampling and testing methods, test samples, review data and report atypical or out-of-specification results under the applicable procedure.
- Engineering and maintenance: confirm equipment, utilities, instruments and facilities are qualified, calibrated and suitable for the intended operation.
- Regulatory Affairs: assess whether the validation approach or proposed change affects a filing, licence commitment or required regulatory notification.
- Qualified release roles: perform any separate certification or batch disposition duties required by the licence and quality system.
4. Definitions
- Critical process parameter (CPP): a process parameter whose variability can affect a critical quality attribute and therefore requires monitoring or control.
- Critical quality attribute (CQA): a physical, chemical, biological or microbiological property that should remain within an appropriate limit to assure product quality.
- Process performance qualification (PPQ): documented evidence that the proposed commercial process, operated with qualified facilities, equipment, personnel and controls, can perform effectively and reproducibly.
- Continued process verification (CPV): ongoing monitoring and review of process performance and product quality during routine manufacture.
- Revalidation: additional or repeated validation work justified by risk, lifecycle monitoring, change or applicable requirements.
5. Process validation procedure
5.1 Establish the strategy and assess risk
The process owner should describe the product, process flow, manufacturing site, equipment train, batch scale, control strategy and intended commercial conditions. Review development knowledge, prior manufacturing experience, technology-transfer information, complaints, deviations, rejected batches and relevant analytical data. Map each unit operation to potential quality risks and identify the controls that address them.
Use a suitable quality risk management method to identify CQAs, potential CPPs, material attributes, in-process controls, sampling points and monitoring needs. Record assumptions, data gaps, risk controls, owners and actions. Revisit the assessment when new knowledge or trends emerge.
5.2 Prepare and approve the protocol
Approve the protocol before executing validation batches or qualification activities. The protocol should be specific enough for trained staff to perform the work and for independent reviewers to evaluate the evidence. Include:
- objective, scope, product and process description, site, equipment and batch scale;
- references to current master manufacturing instructions and linked qualification documents;
- the rationale for the study design, batch number, sampling locations, sample size and test schedule;
- identified CQAs, CPPs, material attributes, in-process controls and data-recording requirements;
- predefined acceptance criteria with a scientific basis and the method for evaluating results;
- responsibilities, training prerequisites, QA oversight and required approvals;
- how deviations, atypical data, missing results, protocol changes and repeat activities will be documented and assessed;
- reporting requirements and the decision process for routine manufacture or further study.
Set acceptance criteria before execution. Criteria should reflect product specifications, process knowledge, analytical capability, variability and regulatory commitments. Do not create or relax limits after seeing results unless an approved change process permits it, with a documented rationale and impact assessment.
5.3 Confirm readiness before execution
Before the study starts, verify that required facilities, utilities, equipment, computerised systems and instruments are qualified or validated for their intended use and within calibration. Confirm that materials have approved status, methods and specifications are current, operators are trained, master instructions are effective, sampling supplies are available, and the protocol has all required approvals.
Document readiness checks and resolve gaps before proceeding. If the protocol permits a justified exception, obtain the required approval in advance and assess its effect on the study.
5.4 Execute process qualification
Manufacture each study batch using the approved instructions and the conditions defined in the protocol. Operators should record actual values, dates, times, equipment identifiers, material lots, interventions, alarms and in-process results when the activity occurs. Reviewers should verify entries for completeness, traceability and good documentation practice.
Collect samples at justified locations and times using approved methods. Assess whether the process operated within intended ranges and whether the control strategy detected or managed variation. Document departures from the protocol promptly; do not omit, replace or repeat data without an approved, traceable rationale.
Evaluate each batch and the study as a whole against the predefined criteria. Use appropriate statistical or trending methods when they suit the data and study objective. Explain exclusions, missing data, atypical results and any differences between batches. A passing result in one test does not by itself establish that the process is controlled.
5.5 Manage deviations and changes
Record every deviation from the approved protocol or manufacturing instructions in the applicable quality system. Assess the cause, scope, data integrity, effect on product quality, affected batches, study validity and need for further work. Define corrective actions and obtain required approvals. Place affected material or batches on hold when the quality system requires it.
Make protocol changes through controlled approval. When a change occurs during execution, document what changed, why, when it took effect and how it affects data already collected. Do not close an unresolved discrepancy through informal annotation.
5.6 Prepare the report and make the approval decision
After data review, the process owner should prepare a report that links evidence to the protocol and validation strategy. Summarise executed activities, batch and equipment details, results, statistical or trend evaluations, deviations, investigations, acceptance-criteria status, limitations and conclusions. Include recommendations, unresolved actions and the proposed routine control strategy.
QA and the designated technical functions should review and approve the report. The conclusion should state whether evidence supports the intended process under the defined conditions, whether additional work is needed, and what controls or monitoring must continue. Document approval before routine use or disposition decisions that depend on validation status.
5.7 Continue verification and control the lifecycle
After qualification, monitor relevant process parameters, in-process controls, yield, rejects, deviations, complaints, out-of-specification results and product quality trends. Define data sources, review frequency, alert or action triggers, responsibilities and escalation pathways. Investigate adverse trends and determine whether corrective action, change control, additional sampling or revalidation is needed.
Route changes to materials, suppliers, equipment, facilities, batch size, process parameters, methods or control strategy through change control. Assess the impact on the validated state, regulatory commitments and ongoing monitoring. The assessment should determine whether no additional validation is needed, targeted verification is sufficient, or a broader study is justified.
6. Process validation checklist
- Validation strategy and risk assessment approved; process knowledge and data gaps reviewed.
- CQAs, CPPs, material attributes, control points and acceptance criteria are defined with rationale.
- Protocol approved before execution; batch design, sampling and data analysis are justified.
- Facilities, utilities, equipment, systems, methods, materials and personnel are ready.
- Study activities and results are recorded contemporaneously and traceably.
- Deviations, atypical data, exclusions and protocol changes are assessed and approved.
- Report links results to criteria, draws a supported conclusion and records required actions.
- CPV, change control, escalation and revalidation triggers are defined for routine manufacture.
7. Frequently asked questions
How many batches are required for process validation?
There is no universal batch count that proves validation in every case. The protocol should justify the number and design based on process knowledge, risk, variability, complexity, scale, regulatory commitments and applicable guidance. Do not present three batches as an automatic rule.
Is retrospective process validation acceptable?
Historical data can support process understanding, risk assessment and lifecycle review. It should not be treated as a universal substitute for prospective process qualification of a new or significantly changed commercial process. Confirm the approach against current requirements and the product-specific regulatory strategy.
What is the difference between qualification and continued verification?
Process qualification provides documented evidence that the proposed process can perform as intended under defined conditions. Continued process verification then monitors routine performance so the site can identify drift, unexpected variability or a need for further action.
When should a process be revalidated?
Assess the need through change control, periodic product and process review, adverse trends, significant deviations, recurring failures, or changes to materials, equipment, facility, scale or process controls. The scope should follow documented risk and applicable regulatory requirements.
8. Regulatory references and related guidance
Consult the current requirements for the product and market. Key references include the FDA’s Process Validation: General Principles and Practices, the European Commission’s EudraLex Volume 4, including Annex 15, and ICH Q10 Pharmaceutical Quality System. For batch record review in the same quality lifecycle, see our batch record review SOP and checklist.
This example is for general information. The organisation’s authorised quality and regulatory personnel must review, approve and adapt it before use as a controlled SOP.