This pharmaceutical stability testing SOP provides an educational framework for planning, conducting, and documenting stability studies for drug substances and drug products. Tailor every protocol to the product, dosage form, container-closure system, target market, marketing authorisation, and applicable quality system. This guide is not an approved regulatory document and does not replace current requirements or professional review.
Key points: Define the study before placing samples; use justified storage conditions and time points; test attributes that can indicate change; document chamber conditions and excursions; and base shelf-life conclusions on the full, reviewed data set.
What ICH Q1A says about stability testing
ICH Q1A(R2), Stability Testing of New Drug Substances and Products, describes a stability data package for registration of new drug substances and products. Its recommendations are applied through the relevant regional regulatory framework. The UK MHRA lists Q1A(R2) among the ICH quality guidelines it has implemented. Confirm the current version and local requirements for each submission or post-approval commitment.
Related guidance may also be relevant: ICH Q1B for photostability, Q1D for bracketing and matrixing, and Q1E for evaluating stability data. These documents have different scopes; do not treat them as a single universal protocol.
Purpose and scope of a stability study SOP
A site procedure should define how the organisation plans studies, controls samples and storage, performs approved tests, reviews results, investigates anomalies, and reports conclusions. It may cover development, registration, commitment, ongoing, in-use, or other studies where applicable. Study design depends on the product and regulatory purpose.
Responsibilities
- Stability coordinator or study owner: prepares the protocol, schedule, sample inventory, and status reports.
- Quality Control: tests samples using approved methods, records original data, and reports atypical, OOS, or OOT results promptly.
- Quality Assurance: reviews and approves controlled documents and evaluates deviations, investigations, and proposed conclusions under site procedures.
- Warehouse or chamber owner: maintains qualified storage equipment, monitoring, alarms, sample access, and excursion records.
- Regulatory or product team: confirms market-specific commitments, approved shelf life, storage statements, and reporting obligations.
Protocol preparation and sample control
Approve a protocol before study execution. It should identify the study objective and applicable guideline; product, batch, strength, manufacturing and packaging details; container-closure configuration; storage condition; sample quantity and pull schedule; test methods and acceptance criteria; responsibilities; data-review steps; and how deviations or excursions will be assessed. Any change to the approved plan should be controlled and justified.
Assign each sample a traceable identifier linked to its batch, package, storage condition, and time point. Record receipt, placement, withdrawal, transfer, and disposition. Protect samples from mix-ups and avoid unplanned exposure during handling. Reconcile scheduled pulls against actual testing and document missed or delayed time points.
ICH Q1A(R2) general-case storage conditions
The following are commonly cited general-case conditions in ICH Q1A(R2) for products intended for room-temperature storage. They are examples, not default instructions for every product. Select conditions for the applicable climatic market, product, packaging, and submission strategy. Refrigerated, frozen, and other special conditions have separate provisions.
| Study condition | General-case example | Use |
|---|---|---|
| Long term | 25°C ± 2°C / 60% RH ± 5% RH, or 30°C ± 2°C / 65% RH ± 5% RH | Chosen according to the applicable market and protocol |
| Intermediate | 30°C ± 2°C / 65% RH ± 5% RH | Used when applicable under the guideline and study design |
| Accelerated | 40°C ± 2°C / 75% RH ± 5% RH | General-case accelerated condition; does not replace long-term data automatically |
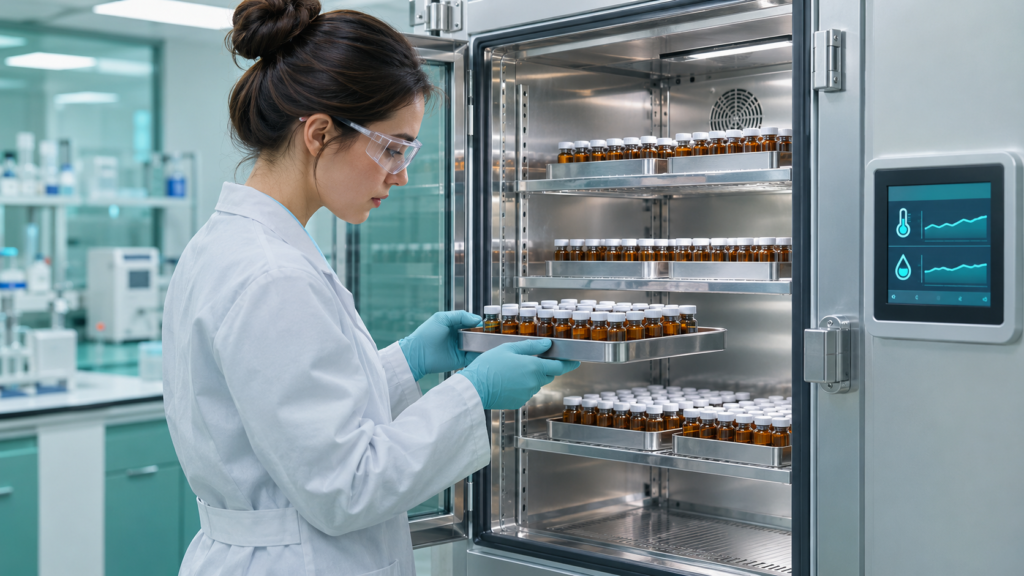
Use qualified chambers capable of maintaining the protocol conditions. Monitor and retain actual temperature and, where relevant, humidity records. Investigate alarms, equipment failures, and excursions under approved procedures; assess and document their potential effect on study interpretation rather than assuming results are invalid or unaffected.
Time points and pull schedule
ICH Q1A(R2) gives general recommendations for formal studies, including frequent testing in the first year of long-term storage and less frequent testing in later years; accelerated studies generally include initial, intermediate, and final time points. The exact schedule must follow the applicable guideline, submission purpose, approved commitment, and protocol. State planned time points explicitly and define how missed pulls are handled.
Select stability-indicating tests for the product
There is no single test panel for every drug substance or product. Select tests that can detect meaningful changes in the product’s quality attributes and are justified by its formulation, dosage form, route, container-closure system, development knowledge, specification, and regulatory commitments. Methods should be suitable for their intended purpose.
- Appearance and physical characteristics, where relevant
- Assay and degradation products or impurities
- Dissolution, disintegration, water content, pH, or functionality tests when applicable
- Microbiological attributes for products where they are relevant
- Container-closure or delivery-system attributes when they can affect product quality
Use the approved protocol and specification to define acceptance criteria. Stability testing supports assessment of quality over time; release tests or generic examples should not be substituted for product-specific shelf-life tests.
Data review, investigations, and shelf-life decisions
Review results against the protocol, specification, historical trends, and study conditions. An OOS result is a result outside an established specification and requires investigation under applicable procedures; it is not automatically a confirmed product failure. An OOT result is an unexpected trend and should be assessed according to the site’s approved procedure. Do not retest, average, exclude, or reinterpret data without a documented scientific basis and required review.
Evaluate the complete data set, including batch variability, method performance, chamber records, deviations, and relevant supporting information. ICH Q1E provides recommendations for evaluating stability data and considering retest periods or shelf lives. Statistical extrapolation is not automatic: it needs scientific justification and must meet the applicable guideline and regional requirements. A proposed expiry or retest period should be supported by the data and approved through the appropriate regulatory and quality process.
Study report and records
- Approved protocol and any controlled amendments
- Sample inventory, storage history, and chamber records
- Raw data, calculations, chromatograms or instrument outputs, and reviewed results
- Deviations, excursions, investigations, and impact assessments
- Trend evaluation, conclusion, proposed shelf-life or retest period, and required approvals
Stability testing checklist
- Confirm current regional guidance, product commitments, and approved specifications.
- Approve the study protocol and test schedule before execution.
- Use representative batches and the intended or justified packaging configuration.
- Verify chamber qualification, monitoring, alarms, and excursion handling.
- Trace each sample through storage, withdrawal, testing, and record review.
- Investigate OOS/OOT results and deviations; document the rationale for conclusions.
- Have the final analysis and proposed shelf-life decision reviewed and approved.
Common stability-study pitfalls
- Applying one storage condition or test panel to dissimilar products or markets
- Using a schedule that does not match the approved protocol or commitment
- Failing to assess chamber excursions and missed time points
- Calling an OOS result a confirmed failure before the required investigation
- Making shelf-life claims from accelerated results alone or unsupported extrapolation
- Leaving out the final packaging configuration or relevant supporting data
Frequently asked questions
Is stability testing mandatory for every pharmaceutical product?
Regulatory submissions and post-approval obligations generally require stability information appropriate to the product and pathway. The exact studies and data package depend on current regional requirements, product characteristics, and approved commitments. Check the applicable rules rather than assuming one universal program.
Are the ICH Q1A(R2) storage conditions suitable for all products?
No. The table above gives general-case examples. Refrigerated, frozen, and other product-specific conditions require the relevant guideline provisions and an approved, justified protocol.
Does accelerated testing establish a product’s shelf life?
Accelerated data support stability evaluation, but do not automatically replace long-term data or establish a shelf life by themselves. Any extrapolation must be justified and meet the applicable regulatory framework.
What is the difference between OOS and OOT in a stability study?
OOS means a result is outside a predefined specification. OOT means a result differs unexpectedly from a relevant trend while it may still meet specification. Both should be handled under approved procedures; an OOS result requires investigation before a conclusion is made.
Can bracketing or matrixing reduce the number of tests?
ICH Q1D describes reduced-design approaches for appropriate cases. Their use needs a scientific rationale showing that the selected design can support the intended conclusions and is acceptable for the relevant submission.
What should a stability study report contain?
Include the approved protocol and amendments, batch and packaging details, storage history, methods, results, trend evaluation, deviations and investigations, conclusions, and required quality and regulatory approvals.
Further reading and official references
For related site guidance, see the finished product testing SOP and batch-release guide and the UK GMP SOP guide.
- ICH Q1A(R2): Stability Testing of New Drug Substances and Products
- ICH Q1E: Evaluation of Stability Data
- ICH Quality Guidelines, including Q1B and Q1D
- MHRA: ICH guidelines implemented in the UK
Educational resource only. It is not an ICH, MHRA, FDA, or EMA approved SOP, nor legal or regulatory advice. Verify current applicable requirements and route any site procedure through your own quality system and approval process.