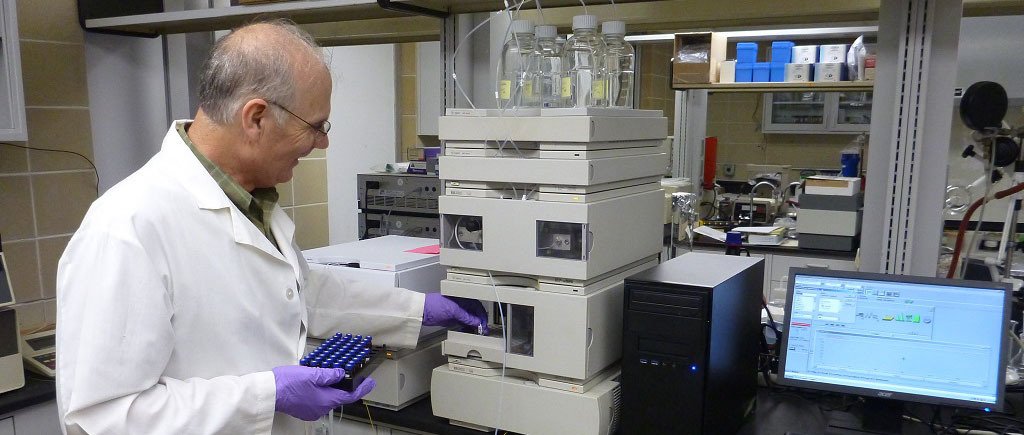
HPLC method development for pharmaceutical analysis is a planned, evidence-based process for creating a chromatographic procedure that is fit for its intended use. This guide explains how pharmaceutical QC and analytical development teams can define the method objective, select chromatographic conditions, prepare samples, evaluate performance and document decisions within the site quality system.
Educational scope: This article is a technical overview, not an approved test method or regulatory advice. Adapt the approach to the product, dosage form, matrix, specification, market commitments and current compendial requirements. Obtain the required scientific and Quality Unit review before implementing a method.
What HPLC method development establishes
A developed method should reliably separate and measure the target analyte or analytes in the relevant sample matrix so that results can support a defined decision. The purpose may be assay, impurities, dissolution, content uniformity, cleaning verification or stability testing. Each application can require different selectivity, range, sensitivity, sample preparation and controls.
Regulatory and quality framework
For US drug CGMP laboratories, 21 CFR 211.160 requires scientifically sound laboratory controls and approved written procedures; § 211.194 addresses complete laboratory records. Other provisions apply according to the test and product, including § 211.165 for testing and release. These regulations are binding where applicable. FDA guidance and ICH guidelines provide recommendations and scientific principles; they do not replace regulations, approved application commitments or applicable compendial requirements.
ICH Q14 describes science- and risk-based analytical procedure development and lifecycle management. ICH Q2(R2) describes validation of analytical procedures. Confirm the current official versions and applicable market expectations. A compendial procedure, a method in an approved filing and a newly developed procedure may follow different verification, validation and change-control pathways.
A practical HPLC method development workflow
1. Define intended use and analytical targets
Document the analyte, product or matrix, reportable range and decision the result will support. Review specifications, known impurities and degradants, sample quantity, solubility, solution stability and relevant regulatory commitments. Set preliminary performance needs from intended use and risk; do not copy generic limits without a product-specific rationale.
2. Select the separation mode, column and detector
Choose chromatographic mode and stationary-phase chemistry based on analyte properties and the separation challenge. Reversed-phase HPLC is common, while HILIC, ion-exchange, normal-phase, chiral or other approaches can be more suitable for particular compounds. Consider column dimensions, particle type, pH compatibility, pressure limits, system dwell volume and laboratory equipment. Select UV, fluorescence, charged aerosol, refractive index or mass spectrometric detection according to the analytical question and method needs; no detector is universally best.
3. Screen mobile-phase and operating conditions
Evaluate variables with a plausible effect on retention, selectivity, peak shape or runtime: organic modifier, buffer identity and concentration, pH, gradient profile, flow, temperature, injection solvent and sample load. Begin with justified screening ranges, then refine influential factors. Change conditions systematically and retain the rationale, actual settings, chromatograms and observations. Design of experiments can help reveal interactions when the design and responses are justified.
4. Develop sample preparation and robustness controls
Demonstrate that extraction, dilution, filtration or other preparation steps recover the analyte and control matrix interference without causing unacceptable degradation or adsorption. Assess solution stability and filter suitability where relevant. Study reasonable variation around critical settings and use the findings to define preparation instructions, system suitability and operating ranges. Confirm the procedure is practical with routine instruments, columns, reagents and trained analysts.
5. Validate, approve and manage the method lifecycle
Prepare an approved protocol with intended use, study design and scientifically justified acceptance criteria before validation. Select relevant characteristics—such as specificity, accuracy, precision, range, linearity, detection or quantitation limits and robustness—based on the method purpose and applicable guidance. Evaluate deviations and limitations, document the conclusion, and obtain required approval before routine use. Manage later changes through the quality system; assess whether verification, partial revalidation or broader work is appropriate.
Development records and routine controls
Maintain traceable records of study plans, reagent and reference-standard details, instrument and column identification, sample preparation, raw data, chromatograms, processing parameters, calculations, deviations, investigations and decisions. The approved procedure should define analyst instructions, system suitability and required records. Use controlled access, review and retention practices under the site’s data-integrity procedures.
For related practical guidance, see our HPLC method troubleshooting guide and our article on current HPLC method development approaches.
Common development pitfalls
- Starting optimisation before defining the intended use and decision.
- Choosing a generic test panel or acceptance limit without product-specific justification.
- Changing several chromatographic variables at once without a documented study plan.
- Ignoring matrix effects, sample stability, filter adsorption or injection-solvent effects.
- Treating a good chromatogram from one trial as proof of robustness or validation.
- Failing to assess instrument capability, method transfer and controlled data processing.
Frequently asked questions
What is the first step in HPLC method development?
Define the intended use, analyte, sample matrix, reportable range and decision. Those factors determine the separation and performance characteristics that matter.
Does every pharmaceutical HPLC method need the same validation tests?
No. Choose validation characteristics and study design for the procedure’s intended purpose, product, matrix, risk, applicable compendial requirements and regulatory commitments. Use current guidance and justified criteria.
Does a reversed-phase column suit every pharmaceutical compound?
No. Reversed phase is widely used, but analyte charge, polarity, solubility and matrix may favour another separation mode or stationary phase. Demonstrate suitable performance for the actual application.
Can automated software approve an HPLC method?
Software can support screening and data analysis, but qualified personnel remain responsible for checking inputs, calculations, model limitations, data integrity and required quality approvals.
Authoritative references
- 21 CFR 211.160 — General requirements for laboratory controls.
- 21 CFR 211.194 — Laboratory records.
- FDA / ICH Q14: Analytical Procedure Development.
- FDA / ICH Q2(R2): Validation of Analytical Procedures.
- FDA: Analytical Procedures and Methods Validation for Drugs and Biologics.
Implementation note: This educational article does not constitute an approved method. Confirm current official requirements and obtain documented site review and approval before using a procedure for GMP testing.