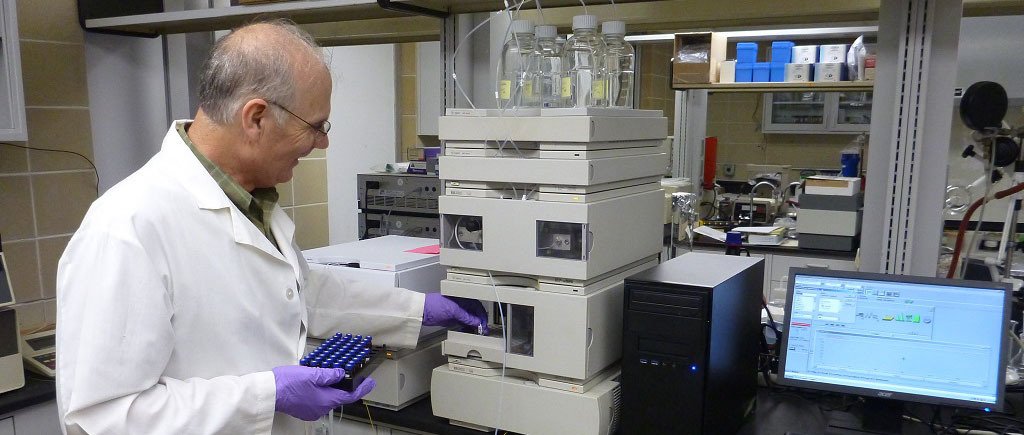
HPLC method optimization is the structured adjustment of chromatographic conditions to achieve a reliable, fit-for-purpose separation. For pharmaceutical analysis, the objective is not simply a shorter run or sharper peak: the procedure must meet its intended performance needs and remain reproducible for the analysts, instruments, samples and lifecycle stage in which it will be used. This guide explains the key HPLC method optimization parameters, a practical screening workflow and the records to retain.
What HPLC method optimization should achieve
Begin with the analytical purpose—such as assay, impurities, dissolution, cleaning verification or a development study—and define the critical analytes, sample matrix, reporting range and required performance. Establish how success will be judged before changing conditions. Depending on intended use, evaluate relevant attributes such as resolution of critical pairs, retention, peak shape, response, precision, run time, carryover and system pressure. Criteria should be justified for the method and product; there is no universal set of HPLC acceptance limits for every application.
Key HPLC method optimization parameters
1. Column chemistry and stationary phase
Choose the separation mode and stationary phase based on analyte properties and the required selectivity. In reversed-phase HPLC, C18 and C8 phases are common starting points, but ligand chemistry, end-capping, pore size, particle characteristics and the sample matrix can change selectivity. Screen an alternative phase when critical compounds co-elute or peak shape remains unsuitable. Check the column manufacturer’s limits for pH, temperature, pressure and solvent compatibility, and document the column identity and dimensions used during development. See our HPLC mobile phase selection guide for related screening considerations.
2. Mobile phase, pH and organic modifier
For ionisable analytes, aqueous-phase pH can affect retention, selectivity and peak shape. Use analyte pKa and stability information to design a sensible pH range, while staying within column and detector constraints. Compare suitable organic modifiers, commonly acetonitrile or methanol in reversed-phase methods, because equal proportions do not necessarily produce equivalent selectivity or pressure. If a buffer or additive is needed, confirm its solubility in the final mixture and compatibility with the detector; non-volatile salts may be unsuitable for LC–MS. Define how the aqueous phase is prepared and how pH is measured so experiments can be reproduced.
3. Isocratic or gradient elution
Use isocratic elution when the analytes have a manageable retention range and the method provides adequate separation. Consider gradient elution for complex samples or a broad range of analyte retention, but evaluate gradient slope, starting and ending composition, hold time and re-equilibration. Confirm that early and late peaks are adequately retained and resolved, and that the system returns to stable baseline and pressure before the next injection.
4. Flow rate, column dimensions and temperature
Flow rate interacts with column diameter, particle size, system dwell volume and pressure. Adjust it within instrument and column limits, then assess separation efficiency and total analysis time rather than relying on speed alone. Column temperature can alter viscosity, retention and selectivity; use controlled, recorded settings and consider analyte stability. When scaling dimensions or transferring a method, reassess flow, gradient delay and system suitability rather than assuming the original conditions transfer unchanged.
5. Detector settings and sample preparation
Select a detector and settings that are appropriate for the analytes and analytical purpose. For UV detection, review wavelength, spectral response and solvent absorbance; for fluorescence or mass spectrometry, consider signal response and additive compatibility. Prepare representative samples using a controlled procedure. Confirm solubility, extraction recovery where relevant, filtration compatibility and diluent strength. A sample diluent much stronger than the initial mobile phase can impair focusing or distort early peaks, so investigate it when peak shape changes.
A practical HPLC method optimization workflow
- Define the target: record intended use, analytes, matrix, critical pairs and fit-for-purpose performance goals.
- Check readiness: confirm instrument status, column condition, reference standards, sample preparation and solvent quality.
- Screen systematically: vary selected factors such as phase, pH, modifier or elution mode in planned experiments; change defined factors and record each condition.
- Optimise critical conditions: assess resolution, retention, peak shape, response, pressure and run time using representative samples and appropriate controls.
- Confirm repeatability and robustness: evaluate relevant sources of variation and verify that the selected conditions remain suitable for their intended use.
- Document and approve: retain protocols, raw data, calculations, chromatograms, rationale, deviations and conclusions; route changes through the site’s change-control and review process.
Pharmaceutical quality, validation and documentation
For US pharmaceutical CGMP work, 21 CFR 211.160 requires scientifically sound laboratory controls and approved written procedures; § 211.194 addresses complete laboratory records. Other provisions may apply according to the product and test, including §§ 211.165 and 211.167. These regulations do not prescribe one universal optimization recipe. Apply the approved analytical procedure, registered commitments, compendial requirements and applicable site quality system.
FDA’s ICH Q14 Analytical Procedure Development guidance and ICH Q2(R2) Validation of Analytical Procedures guidance describe science-based development and validation principles. FDA guidance represents the Agency’s current thinking and is generally nonbinding; it does not replace applicable regulations or product-specific commitments. Validation, verification or other lifecycle work should be selected for the procedure’s intended use and relevant change.
In a regulated laboratory, retain enough information for another qualified analyst to understand what was tested and why: experimental plan, reagent and mobile-phase preparation, instrument and column identifiers, sequences, processing method, original data, calculations, observations, investigation records and approval decisions. Apply data-integrity controls and do not discard or selectively report results that do not support the preferred condition.
Common HPLC optimization mistakes
- Changing multiple variables at once, making it difficult to interpret cause and effect.
- Choosing a condition solely because it shortens the run, without checking resolution, carryover, pressure or repeatability.
- Ignoring pH, sample diluent, buffer precipitation, column limits or detector compatibility.
- Using non-representative samples or standards that do not challenge the separation.
- Treating development experiments as validation, or implementing a changed method before required review and approval.
- Keeping only the final chromatogram instead of the study plan, original records, unsuccessful trials and scientific rationale.
Frequently asked questions
What are the most important HPLC method optimization parameters?
The most influential parameters often include stationary phase, mobile-phase composition and pH, elution mode, flow rate, column dimensions, temperature, detector settings and sample preparation. Which factors matter most depends on the analytes, matrix and method purpose.
How do I improve resolution in HPLC?
First identify which peaks are critical and what is limiting separation. Screen selectivity changes such as stationary phase, pH or organic modifier, then consider gradient, flow or temperature within operating limits. Change factors systematically and confirm performance with representative samples.
Does every optimized HPLC method need validation?
Not every change has the same validation impact. Determine appropriate validation, verification or other documented assessment from intended use, method lifecycle stage, the nature of the change and applicable requirements. Use the site’s approved procedures and relevant guidance.
What should be recorded during HPLC method development?
Record the objective, study plan, exact chromatographic conditions, column and instrument details, sample and standard preparation, raw data, processing, results, deviations, decisions and rationale. The documentation should support traceability and reproducibility.
Conclusion
Effective HPLC method optimization uses planned experiments to balance selectivity, resolution, robustness, analysis time and instrument constraints. Define the method’s purpose first, investigate the parameters that matter for the separation, and document the evidence behind the final conditions. For pharmaceutical use, assess the procedure through the applicable quality system and obtain required technical review and approval before routine implementation. For related guidance, read our HPLC troubleshooting guide.